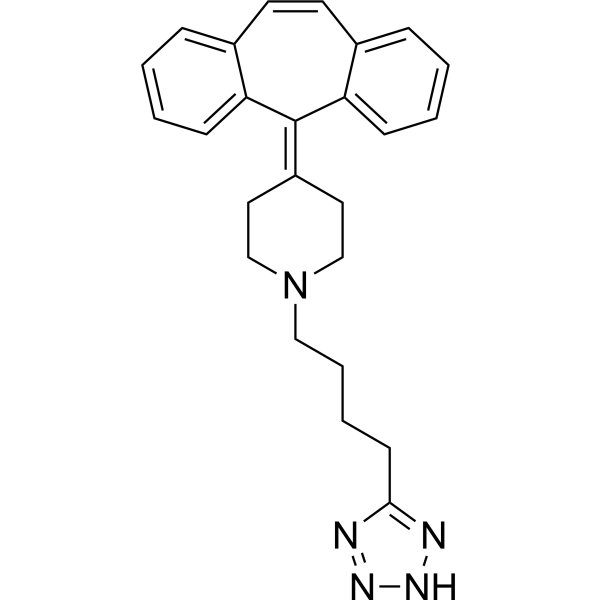
AT-56 chemical structure
Chemical structureResearch use only
CAT. NO. DC7364Featured

AT-56

Pack Sizes & Prices

TotalUS$550
We match the best price and quality on market.
Product overview

Description & Application

AT-56 is a potent, selective and orally active inhibitor of lipocalin-type prostaglandin D synthase (L-PGDS), with an IC50 of 95 μM and Ki of 75 μM. AT-56 could selectively suppress the drowsiness or pain reaction mediated by L-PGDS-catalyzed PGD2[1].

ParameterValue
Chemical NameAT 56
CAS No.162640-98-4
SynonymsAT 56;4-(5H-Dibenzo[a,d]cyclohepten-5-ylidene)-1-[4-(2H-tetrazol-5-yl)butyl]-piperidine;AT-56;4-(DIBENZO[1,2-A:1',2'-E][7]ANNULEN-11-YLIDENE)-1-[4-(2H-TETRAZOL-5-YL)BUTYL]PIPERIDINE;4-Dibenzo[a,d]cyclohepten-5-ylidene-1-[4-(2H-tetrazol-5-yl)-butyl]-piperidine;4-(5H-Dibenzo[a,d]cyclohepten-5-ylidene)-1-[4-(1H-tetrazol-5-yl)butyl]piperidine
FormulaC25H27N5
M.Wt397.51538
SMILESC1=CC=C2C(=C1)C=CC3=CC=CC=C3C2=C4CCN(CCCCC5=NNN=N5)CC4
Purity>98%
Storage2 years -20°C Powder, 2 weeks 4°C in DMSO, 6 months -80°C in DMSO

Biological Activity

AT-56 is a potent, selective and orally active inhibitor of lipocalin-type prostaglandin D synthase (L-PGDS), with an IC50 of 95 μM and Ki of 75 μM. AT-56 could selectively suppress the drowsiness or pain reaction mediated by L-PGDS-catalyzed PGD2[1].

In Vitro

AT-56 (1-30 μM; 10 minutes) dose-dependently inhibits the production of PGD2 in L-PGDS-expressing human medulloblastoma TE-671 cells with an IC50 of about 3 μM[1].

In Vivo

AT-56 ( 1-30 mg/kg; p.o.) suppresses the PGD2 production in the stab-wounded brain[1]. AT-56 (1-10 mg/kg; p.o.) suppresses the L-PGDS-mediated allergic airway inflammation in mice[1]. AT-56 (10 mg/kg; p.o.) exhibits Cmax (2.15 μg/ml), half-life (1.71 h) and high oral bioavailability (82%)[1]. Animal Model: H-PGDS KO mice (14-16weeks, 25-30 g, C57BL/6 strain) with a stab wound brain injury[1] Dosage: 0, 1, 3, 10, 30 mg/kg Administration: P.o. 1 h before the stab wound injury Result: Inhibited the L-PGDS reaction in the brain. Decreased the total amount of PGD2 in the brain to 40% with 30 mg/kg AT-56. Animal Model: Human L-PGDS-overexpressing TG mice (males, 14-16 weeks, 25-30 g)[1] Dosage:  0, 1, 10 mg/kg Administration: P.o. 1 h before and 24 h after the antigen exposure Result: Prevented the eosinophil infiltration by inhibiting transgened human L-PGDS. Animal Model: Male C57BL/6 mice (7 weeks, 22-26 g)[1] Dosage: 10 mg/kg for p.o. and 2 mg/kg for i.v. (Pharmacokinetic Analysis) Administration: P.o. and i.v. administration Result: Oral bioavailability (82%); Cmax (2.15 μg/ml); T1/2 (1.71 h, p.o.); T1/2 (2.35 h, i.v.).

References & source

DC Chemicals original product information

Published experimental findings depend on the model and assay conditions. They do not guarantee equivalent performance in every application. For laboratory research use only.
Product documentation

Refer to the certificate supplied with your lot for batch-specific release results.

Handling guide

2 years -20°C Powder, 2 weeks 4°C in DMSO, 6 months -80°C in DMSO

Solution calculator

Related research products

GW-627368X chemical structureDC8266

GW-627368X

GW627368(GW627368X) is a novel, potent and selective competitive antagonist of prostanoid EP4 receptor(Ki= 100 nM) with additional human TP receptor affinity(Ki= 150 nM).IC50 value: Target: EP4 antagonistin vitro: At…

View activity & prices
E-7046 chemical structureDC10563

E-7046

E7046 is an orally bioavailable and specific EP4 antagonist, with IC50 of 13.5 nM and Ki of 23.14 nM, exhibiting anti-tumor activities.

View activity & prices
TG6-10-1 chemical structureDC8348

TG6-10-1

TG6-10-1 is an EP2 antagonist, shows low-nanomolar antagonist activity against only EP2, >300-fold selectivity over human EP3, EP4, and IP receptors, 100-fold selectivity over EP1 receptors.

View activity & prices
Fevipiprant chemical structureDC11278

Fevipiprant

Fevipiprant(QAW039) is a selective, potent, reversible competitive CRTh2 antagonist with an in vitro dissociation constant KD value of 1.1nM at the CRTh2 receptor and an IC50 value of 0.44 nM for inhibition of…

View activity & prices
AZD1981 chemical structureDC7572

AZD1981

AZD1981 is a potent and selective CRTh2 antagonist; displaces radio-labelled PGD2 from human recombinant DP2 with high potency (pIC50 = 8.4).

View activity & prices
Dinoprost chemical structureDC9863

Dinoprost

Dinoprost(Prostaglandin F2α) is a naturally occurring prostaglandin used in medicine to induce labor and as an abortifacient.

View activity & prices
TG8-260 chemical structureDC70837

TG8-260

TG8-260 is a second-generation, potent, selective EP2 antagonist with Schild KB of 13.2 nM.TG8-260 displays 500-fold selectivity to EP2 against other prostanoid receptors.TG8-260 has a plasma half-life of 2.14 h (PO)…

View activity & prices

Cart

Request quotation