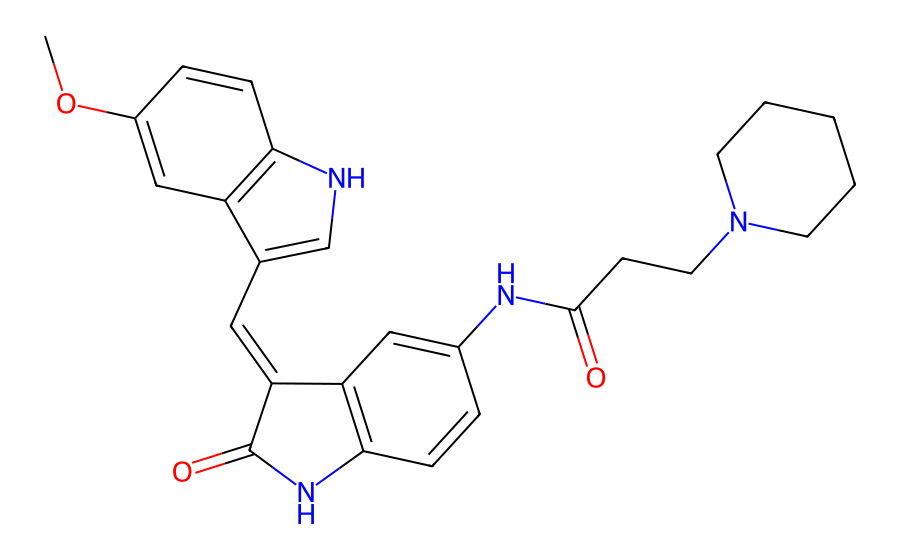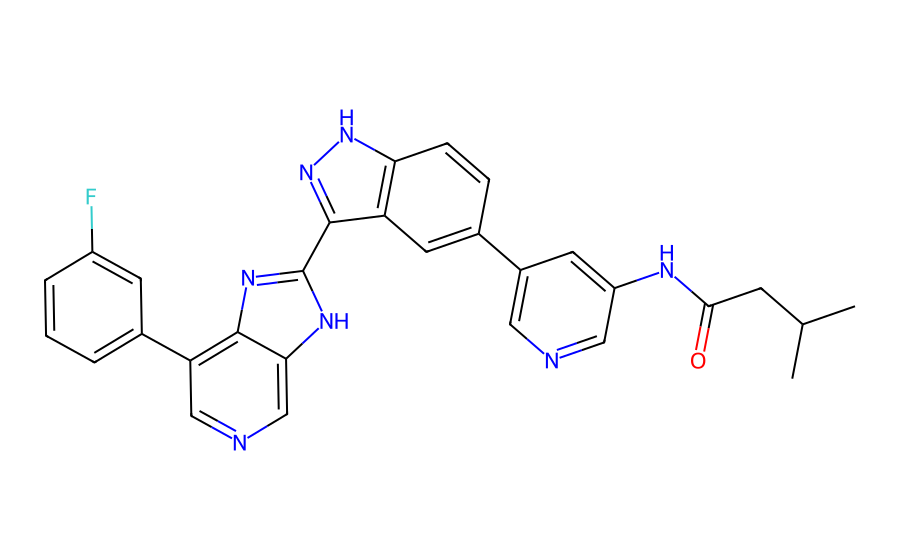
Biological Activity
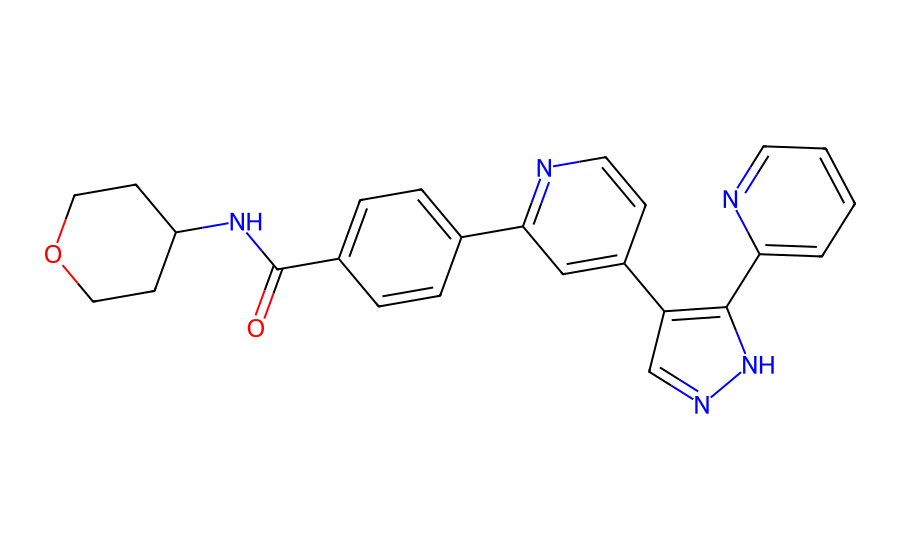
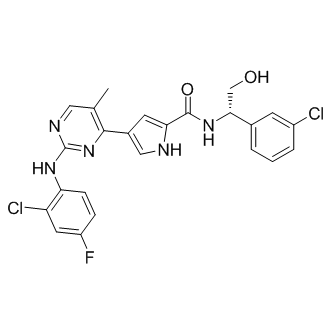
VX-11e is a potent, selective, and orally bioavailable inhibitor of ERK with Ki < 2 nM.
In Vitro
VX-11e is active in the HT29 cell proliferation assay (IC50=48 nM)[1].
In Vivo
VX-11e is orally bioavailable in both rat and mice[1]. VX-11e (50 mg/kg, p.o.) results in robust inhibition of pRSK, and inhibits tumor growth in NSG mice bearing human melanoma RPDX tumors. VX-11e with BKM120 significantly improves tumor growth inhibition[2].
Target
ERK2:2 nM (Ki)
GSK3:395 (Ki)
AURA:540 (Ki)
CDK2:852 (Ki)
Kinase Assay
Compounds are assayed for the inhibition of ERK2 by a spectophotometric coupled-enzyme assay. In this assay, a fixed concentration of activated ERK2 (10 nM) is incubated with various concentrations of the compounds in DMSO (2.5%) for 10 min. at 30°C in 0.1 mol/L HEPES buffer, pH=7.5, containing 10 mM MgCl2, 2.5 mM phosphoenolpyruvate, 200 μM NADH, 150 μg/mL pyruvate kinase, 50 μg/mL lactate dehydrogenase and 200 μM erktide peptide. The reaction is initiated by the addition of 65 μM ATP. The rate of decrease of absorbance at 340 nM is monitored.
Cell Assay
Cell proliferation is measured by 3H-thymidine incorporation. The cells are plated at a concentration of 10,000 cells/well in a 96-well plate using growth media, RPMI 1640 containing 10% FBS. Serially diluted compounds are added. The cells and compounds are incubated for 48 hours at 37°C incubator. After 48 hours, 0.4 μCi of 3H-thymidine is added to each wells for 8 hours and returned to the 37°C incubator. The cells are harvested using a Tomtec 96-well cell harvester and the CPM is determined using the Wallac 1205 BETAPLATE liquid scintillation counter.
Animal Administration
Human melanoma RPDX tumors are expanded in vivo using NSG mice prior to the therapy experiments. Pooled tumor chunks banked from early mouse passages are implanted into 50 NSG mice (1:10 expansion). These tumors are harvested when reaching the maximum volume allowed on the protocol (1,000 mm3), digested, and banked as live cells. The larger part of this stock is retained as a master bank, and the other part is implanted at a 1:5 ratio into NSG mice to use in the therapy experiments. The expansion phase is under continuous drug pressure with PLX4720 200 ppm chemical additive diet at approximately clinical plasma levels. The plasma levels of PLX4720 (103.7 μg/mL ±3.2 after 7 days) are similar to steady-state levels in patients treated with vemurafenib 960 mg twice a day (130.6 μg/mL±71.78). When tumors have reached 200 mm3 per caliper measurement, animals are randomized into treatment groups followed by a 3-day ishout phase. Tumor size is assessed twice weekly per caliper measurement. Mice are sacrificed after two weeks of treatment or when necessary for animal welfare. Dosing is prolonged when tumor control is achieved as indicated. Tumor tissue is conserved in formalin (for FFPE) and snap-frozen in liquid N2 for protein extraction. Treatment groups are sacrificed 4 hours after last dose.
References & source
[1]. Aronov, Alex M., et al. Structure-Guided Design of Potent and Selective Pyrimidylpyrrole Inhibitors of Extracellular Signal-Regulated Kinase (ERK) Using Conformational Control. Journal of Medicinal Chemistry (2009), 52(20), 6362-6368.
[2]. Krepler C, et al. Personalized Preclinical Trials in BRAF Inhibitor-Resistant Patient-Derived Xenograft Models Identify Second-Line Combination Therapies. Clin Cancer Res. 2016 Apr 1;22(7):1592-602.
Published experimental findings depend on the model and assay conditions. They do not guarantee equivalent performance in every application. For laboratory research use only.